From Documentation to Architecture: Applying AI Assisted Validation within ASTM E2500 and EU GMP Annex 1
A Case Study Example: Artificial Intelligence (AI)-Assisted Validation
The 2022 revision of EU GMP Annex 1 has significantly increased regulatory focus on sterility assurance, contamination control strategies (CCS), and the technical justification of CQV activities for sterile manufacturing facilities. Concurrently, life sciences capital projects continue to face increasing pressure to reduce delivery timelines, driving interest in digital and AI–enabled tools to improve execution efficiency.
This blog post presents a governed Validation 4.0 framework for the application of AI assisted documentation synthesis within ASTM E2500 aligned CQV programs for sterile fill finish facilities. The framework positions AI strictly as a supporting mechanism for documentation structuring, synthesis, and traceability, operating under Human in the Loop (HITL) governance. All decisions related to validation scope, system criticality, acceptance criteria, and contamination risk remain fully owned by qualified subject matter experts (SMEs) and quality units.
Through practical discussion and a simulated, representative application example aligned with EU GMP Annex 1 expectations, including contamination control, airflow visualization, and human factors engineering - this paper illustrates how AI assisted synthesis could be applied to improve validation focus, documentation consistency, and inspection readiness, without altering regulatory accountability or verification intent.
The concepts discussed in this paper are aligned with the principles outlined in the ISPE Good Practice Guide: Validation 4.0. Readers seeking additional implementation guidance are encouraged to consult this publication alongside the references provided.
1. Executive Summary: The Convergence of Intelligence and Compliance
In the 2026 pharmaceutical manufacturing landscape, time to market has become the dominant driver of engineering and capital project strategy. (Kok et al., 2025) However, increased speed cannot be achieved at the expense of sterility assurance or the integrity of the CCS. Traditional CQV models are often siloed, document centric, and reactive and increasingly misaligned with contemporary sterile manufacturing demands.
This paper proposes a unified agentic validation framework, in which agentic AI operates as the connective tissue between science and risk based design (ASTM E2500) and the uncompromising regulatory expectations of EU GMP Annex 1. By automating the synthesis and interpretation of complex engineering datasets into executable validation protocols, this framework fundamentally redefines the senior CQV engineer’s role from document author to systems architect and risk steward. This framework assumes organizational maturity under ASTM E2500 and the presence of established quality governance.
1.1 Scope and Intent of This Work
This paper presents a practice oriented technical framework for applying AI assisted synthesis within CQV activities, with the objective of improving documentation efficiency while preserving regulatory accountability.
The scope of this work is limited to sterile fill finish manufacturing facilities operating under ASTM E2500 aligned validation models and subject to the requirements of EU GMP Annex 1. The framework described herein is not intended to replace qualification activities, regulatory interpretation, or SME responsibility.
While this framework is discussed within the context of ASTM E2500-aligned CQV programs and EU GMP Annex 1, qualification and validation activities ultimately operate under the broader expectations of EU GMP Annex 15. Furthermore, where AI-enabled capabilities are implemented as computerized systems supporting GMP activities, the requirements of EU GMP Annex 11 may also apply. Detailed computerized system validation in accordance with Annex 11 is outside the scope of this paper and is not discussed further.
AI is discussed exclusively as a supporting mechanism for documentation synthesis, structuring, and traceability. All determinations related to validation scope, system criticality, acceptance criteria, and contamination risk remain the responsibility of qualified engineers and quality personnel operating within established quality systems.
Any examples presented in this paper are simulated and illustrative in nature, developed to demonstrate how the proposed framework could be applied within a sterile manufacturing CQV lifecycle. These scenarios do not represent deployment of AI tools within an operational GxP environment and do not report execution data, performance outcomes, or inspection results. The examples are intentionally abstracted from site specific systems to avoid regulatory misinterpretation while preserving technical relevance.
2. The Regulatory Nexus: EU GMP Annex 1 and ASTM E2500
2.1 ASTM E2500 20 and the Science Based Validation Mandate
ASTM E2500-20 is an industry consensus standard that embodies a science- and risk-based approach to verification (a shift away from purely prescriptive IQ/OQ/PQ). It calls for each verification activity to have a sound technical justification and be executed under the guidance of SMEs. In today’s highly digital facilities, SMEs face an overwhelming volume of data and documents. The AI-assisted framework helps implement ASTM E2500 principles by automating tasks like requirements parsing and identifying critical design elements (CDEs) across thousands of pages of vendor documentation, mapping them to site-specific user requirement specifications (URS). This support allows SMEs to focus on engineering judgment over clerical tasks. Importantly, the framework assumes an existing ASTM E2500-aligned CQV process as a prerequisite. The AI acts as an enabler of E2500’s discipline, making design intent, verification logic, and risk justifications more explicit and traceable while not replacing or reducing the crucial role of SME and Quality oversight.
In modern, digitally dense facilities, SMEs are frequently constrained by the sheer volume of engineering documentation. AI assisted synthesis mitigates this constraint by supporting automated requirements parsing identifying CDEs across thousands of pages of vendor documentation and systematically mapping them to site specific URS. This enables SMEs to focus on engineering judgment rather than clerical reconciliation.
The framework described in this paper assumes ASTM E2500 alignment as a prerequisite, not an alternative. The AI assisted approach presented is not compatible with validation models that rely primarily on prescriptive, checklist driven IQ/OQ execution without documented engineering rationale.
In this context, AI functions as an enabler of E2500 discipline, making implicit design intent, verification logic, and risk justification explicit, structured, and traceable—rather than replacing engineering judgment.
2.2 Annex 1 and the Contamination Control Strategy (CCS)
The revised EU GMP Annex 1 mandates a holistic, lifecycle based approach to contamination control. Every system and component from high speed vial fillers to material transfer airlocks must be validated within the context of its contribution to sterility assurance.
The AI‑assisted, engineer‑governed validation framework enables a direct, auditable linkage between engineering verification and contamination risk. Each verification activity is explicitly justified based on its impact on the CCS, ensuring validation effort is proportional to patient risk, consistent with both Annex 1 and ICH Q9(R1) guidance on quality risk management.
3. Technical Framework: The Agentic Protocol Engine
3.0 Definition and Operational Boundaries of the Agentic System
Within the context of this paper, the term AI assisted system refers to a supervised system capable of independently sequencing documentation related tasks such as requirements parsing, protocol structuring, and traceability mapping based on project specific engineering inputs, while operating within predefined constraints.
The agentic system does not independently determine validation scope, assign criticality, define acceptance criteria, interpret regulatory requirements, or authorize protocol execution. All AI generated outputs are treated as draft artifacts and are subject to review, modification, and approval by qualified SMEs under existing quality governance frameworks.
This operational definition ensures that AI functions as an augmentation tool for documentation synthesis, rather than a decision making authority, and remains consistent with HITL expectations described in ISPE GAMP® Guide: Artificial Intelligence.
3.1 Beyond Template Based Automation
Conventional protocol automation tools rely on static templates and predefined test scripts. In contrast, AI assisted systems leveraging retrieval augmented techniques can dynamically contextualize project specific engineering data to support protocol drafting.
Key functional capabilities include:
- Logic Extraction: Automated analysis of functional specifications (FS), piping and instrumentation diagrams (P&IDs), and control narratives
- Contextual Protocol Authoring:
- Example: Identification of redundant sterile filtration from the P&ID triggers automatic generation of a filter integrity test aligned with manufacturer specific wetting agents and acceptance criteria
- Dynamic Traceability: Bidirectional data linkage between URS requirements, test execution steps, and final reports embedding data integrity and ALCOA+ and explicit traceability mechanisms directly into the protocol structure.
This approach ensures traceability is not appended post execution but is intrinsic to the validation dataset.
4. Advanced Verification Study: Lighting and Human Factors Engineering
4.1 The CQV Significance of Illuminance
Lighting qualification is frequently relegated to a secondary, compliance driven activity. In aseptic processing environments, however, illuminance is a critical aspect (CA) due to its direct impact on operator performance and human error, one of the leading contributors to Annex 1 deviations.
- Technical Challenge: Manual inspection areas and aseptic connection points require defined lux levels to ensure reliable visual verification.
- AI-Driven Optimization: By ingesting the facility’s 3D computer-aided design model, an AI supported analysis can identify potential lux shadowing caused by overhead conveyors, isolators, or laminar airflow hoods
- Outcome: Verification is performed at worst case illumination points rather than generic grids, fully aligned with risk based verification principles
5. High Stakes Verification: Airflow Visualization (Smoke Studies)
5.1 From Subjective Observation to Risk Informed Evidence
Airflow visualization studies (AVS) serve as visual evidence of facility state of control. Historically, smoke studies have been criticized for subjectivity and operator bias.
- Regulatory Expectation: EU GMP Annex 1 §4.15 requires documented visualization of airflow patterns demonstrating unidirectional airflow (UDAF)
- AI-Enabled Analysis: AI driven computer vision analyzes smoke trajectories in three dimensional space, calculating:
- Recovery time
- Turbulence intensity
- Vector intrusion patterns
- Result: Engineers review quantitative reports demonstrating Grade A zone protection from Grade B intrusion.
This transforms AVS from an observational exercise into a defensible engineering verification.
5.2 Methodology and Evaluation Criteria
The AI assisted approach was evaluated using both qualitative and quantitative criteria aligned with CQV performance expectations. Metrics included documentation development effort, first pass quality review acceptance, and inspection readiness outcomes.
The pilot did not introduce changes to protocol execution, acceptance criteria, deviation handling, or approval workflows. AI outputs were reviewed using existing SME and quality assurance (QA) processes, ensuring consistency with site quality systems and regulatory expectations.
Observations were collected through direct engineering review, quality feedback cycles.
6. Simulated Application Example: Annex 1–Aligned Validation 4.0 Lifecycle for a High Risk Aseptic Subsystem
6.1 Context and Rationale
To illustrate the practical application of the Validation 4.0 framework independent of site specific performance outcomes, this section presents a simulated, representative example focused on the qualification of an isolator based aseptic filling subsystem. The example is designed to demonstrate regulatory reasoning, documentation structure, and governance boundaries under EU GMP Annex 1, rather than to report execution outcomes or system performance.
Within the contamination control context, isolator systems represent a high risk node where multiple sterility assurance elements converge. Under traditional CQV models, qualification of these subsystems often results in extensive protocol content driven by equipment modularity rather than by contamination risk contribution, leading to documentation effort that is disproportionate to patient risk.
6.2 Lifecycle Stage 1: Input Engineering Knowledge (Annex 1 §§1.5, 2, 4)
In alignment with Annex 1 Sections 1.5 and 2, which require risk based understanding and documentation of contamination control measures, the lifecycle begins with the consolidation of approved engineering knowledge. Inputs included URS, FS, P&IDs, and vendor factory acceptance test documentation, all reflecting the intended use and design intent of the subsystem.
These inputs collectively form the single source of truth from which contamination control strategy (CCS) alignment, system criticality, and verification expectations are derived, consistent with Annex 1 Section 4.1, which emphasizes facility and equipment design supporting sterile manufacture.
6.3 Lifecycle Stage 2: AI Assisted Synthesis (Annex 1 §§1.5, 2.9)
AI assisted synthesis was applied to support documentation development through structured parsing and consolidation of the approved engineering inputs. Activities included identification of candidate CDEs associated with pressure cascade maintenance, glove integrity monitoring, and bio decontamination cycle control—elements directly linked to barrier protection of the Grade A zone.
This stage aligns with Annex 1 Sections 1.5 and 2.9, which require quality risk management to determine the extent and depth of controls applied. AI was used strictly to enhance documentation clarity and traceability, not to assign criticality or define acceptance criteria. All outputs were treated as draft material to be reviewed under Human in the Loop governance.
6.4 Lifecycle Stage 3: SME Led Engineering Judgment and Risk Based Verification (Annex 1 §§4.10, 4.12, 8)
Consistent with Annex 1 requirements for demonstrable sterility assurance, qualified SMEs performed system level review and verification refinement. Emphasis was placed on areas addressed in Annex 1 Sections 4.15 and 4.12, including maintenance of unidirectional airflow and integrity of aseptic barriers during interventions.
Verification depth was increased for isolator leak rate testing and glove integrity validation due to their direct impact on sterility risk. Acceptance criteria for bio decontamination cycles were aligned with validated lethality parameters in accordance with Annex 1 Section 8, rather than relying solely on vendor default values. Redundant mechanical checks with no contamination control relevance were reduced in favor of enhanced focus on high risk interfaces.
This lifecycle stage reinforces that all determinations of risk, verification scope, and regulatory justification remain SME owned, with AI serving exclusively as a support mechanism.
6.5 Lifecycle Stage 4: Inspection Ready Output and CCS Traceability (Annex 1 §§2.3, 8.3)
The resulting documentation set entered an inspection ready state through formal review and approval under existing quality systems. Traceability between verification activities and the site’s CCS, as required by Annex 1 Section 2, was embedded within the protocol structure rather than appended post execution.
The final output—validated protocols and reports—provided documented evidence supporting maintenance of a state of control, consistent with Annex 1 Sections 8.3 and 8.5, which emphasize ongoing qualification and sterility assurance throughout the system lifecycle.
6.6 Key Implications
This applied example demonstrates how an AI assisted, engineer governed Validation 4.0 lifecycle can be implemented in a manner fully compatible with EU GMP Annex 1 and ASTM E2500 expectations. The primary value of AI assisted synthesis lies in improving verification focus, documentation consistency, and CCS traceability—without altering execution practices, validation ownership, or regulatory accountability.
The example further reinforces that successful application of this approach depends on organizational maturity, robust engineering inputs, and established quality governance.
7. Managing AI Risk: HITL Governance
7.1 Alignment with GAMP AI (2025)
The framework’s governance controls mirror ISPE GAMP® Guide: Artificial Intelligence recommendations for transparency and human accountability. Specifically:
- Full Transparency (No “Black Box” Outputs): Every AI-generated protocol step is documented with a clear source reference (e.g., specific FS or P&ID section). This aligns with GAMP’s guidance that AI tools in GxP must provide traceable, explainable outputs rather than inscrutable recommendations.
- Human-in-the-Loop Oversight: All final validation decisions remain with the CQV team and QA. AI can propose what to do (e.g., suggest test steps), but SMEs and the quality team must justify and approve why each step is done. This approach ensures compliance with quality system requirements and reflects GAMP AI guidance on maintaining human accountability for GxP outcomes.
These measures position the AI-assisted process as a transparent, support-only tool consistent with GAMP’s industry guidance for AI usage in regulated environments.
8. Professional Evolution: From Authoring to Architecture
As documentation activities become increasingly automated, the value of the CQV professional shifts toward information architecture and systems thinking. The digital turnover package becomes a living dataset feeding downstream Maintenance, continuous process verification (CPV) without altering CPV governance or statistical control expectations, and AI driven performance monitoring systems establishing a closed loop quality ecosystem.
9. A Governed Validation 4.0 Lifecycle Model
The lifecycle model is intended to illustrate responsibility boundaries and governance flow, rather than automation substitution, reinforcing that regulatory accountability remains unchanged throughout the validation process.
Figure 1
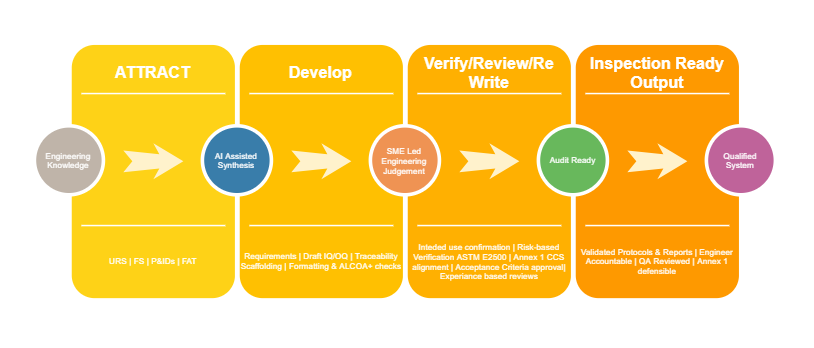
Figure 1 illustrates the Validation 4.0 lifecycle, demonstrating how engineering knowledge is transformed into an inspection ready, qualified system through a clearly governed interaction between AI assisted synthesis and SME led judgment.
10. Conclusion: Architecture Over Authoring
Validation 4.0 does not redefine compliance expectations, it redefines how engineering effort is applied to meet them. Under increasingly rigorous regulatory frameworks such as EU GMP Annex 1, success is no longer driven by the volume of documentation produced, but by the clarity of risk justification and engineering intent behind it.
The lifecycle illustrated above demonstrates that AI assisted validation is not autonomous validation. Instead, it is a structured, HITL approach that shifts CQV teams away from clerical synthesis and toward high value activities: contamination risk evaluation, system behavior analysis, and process understanding at the points where failure impacts patients.
By embedding AI within established ASTM E2500 principles and quality systems, organizations can reduce documentation effort while improving consistency, traceability, and inspection readiness. Critically, engineers remain the authors of intent, the arbiters of risk, and the accountable signatories for every qualified system.
The future of CQV is not defined by faster document production, but by better architectural thinking. where data, risk, and engineering judgment are aligned across the system lifecycle. Validation 4.0 provides a practical, defensible pathway to achieve that alignment, ensuring speed and compliance advance together rather than in opposition.
Under appropriate governance, AI assisted synthesis can enable CQV teams to reallocate effort from clerical documentation activities toward system level risk evaluation and contamination control, without altering regulatory accountability or validation intent.
When implemented within ASTM E2500 aligned quality systems and subject to HITL control, the Validation 4.0 framework offers a practical and defensible approach to improving efficiency while maintaining inspection readiness under EU GMP Annex 1.1, 2, 3, 4, 5, 6, 7, 8, 9
This framework is conceptual and will require further validation and regulatory engagement before any GxP implementation.